Top eQMS Requirements for Biotech Companies

Biotech is not a steady-state industry. A company can move from a handful of researchers to a multisite commercial operation in a matter of years, sometimes months, and the quality system has to keep up at every stage. For teams choosing or upgrading an eQMS for biotech, the stakes are higher than picking software that checks regulatory boxes. The system has to hold together through headcount surges, shifting processes, and the transition from early-stage development to commercial manufacturing, all without losing the data integrity and audit readiness that regulators expect from day one.
This post walks through the eQMS requirements that matter most to biotech companies, covering scalability as the business grows, validation of the system itself, change management across processes and documents, and data integrity controls that stand up to scrutiny. Each of these areas has specific, concrete implications for what a quality management system needs to do, and for what a growing biotech should look for before committing to one.
Why Biotech Companies Have Distinct eQMS Needs
Rapid Growth Changes the Requirements Themselves
A biotech company’s quality needs at 20 employees look nothing like its needs at 200. Early on, a small team might manage document control and training records in spreadsheets or shared drives. That approach breaks down quickly once multiple departments, contract manufacturers, and new product lines enter the picture. The quality system a company adopts needs to absorb that growth without a disruptive platform change every time the org chart shifts, because replacing a QMS mid-scale-up introduces its own validation burden and operational risk.
Frequent Process Change Is the Norm, Not the Exception
Biotech companies iterate. Manufacturing processes get refined as a product moves through development, analytical methods get updated, and SOPs evolve as lessons from audits, deviations, and tech transfers get incorporated. A quality system built for a stable, slow-moving manufacturer will struggle with this pace. Biotech companies need a QMS that treats frequent, well-documented change as routine business, not as an exception that requires a workaround.
The Early-Stage to Commercial-Stage Leap
Few transitions put more pressure on a quality system than the move from clinical-stage to commercial-stage operations. Documentation expectations, regulatory scrutiny, and the volume of quality records all increase substantially at that point. A company that chose a QMS suited only to an early-stage pace, or relied on manual processes, often finds itself replatforming right when it can least afford the distraction of a system migration. Planning for that transition early, even before it’s imminent, is one of the more overlooked aspects of GxP compliance strategy.
What a Scalable, Growth-Ready eQMS Needs to Support
A scalable QMS is not just software that happens to work at a larger company. It is a system designed from the outset to flex with headcount, site count, product count, and regulatory complexity. For a growing biotech, that means the eQMS should support the following.
- Configurable workflows that don’t require custom code. As roles, approval chains, and departments change, workflow logic should be adjustable through configuration rather than a development project.
- Multisite and multientity support. Companies that add contract manufacturers, additional facilities, or international sites need a QMS that can manage distinct but connected quality processes across locations.
- Modular rollout. A system that supports starting with core modules (document control, training, CAPA) and adding others (supplier quality, audit management, complaint handling) as the business matures avoids forcing an all-or-nothing implementation before the company is ready for it.
- User and role management that scales. Onboarding new employees, contractors, and departments should be straightforward, with access and training requirements tied to role rather than manually reconfigured each time.
- Data structures that hold up under volume. As the number of quality records, batches, and products grows, the system needs to maintain performance and traceability rather than becoming harder to search and report on.
A quality management system that can’t flex in these ways tends to generate workarounds, and workarounds are exactly what erode data integrity and audit readiness over time.
Validation Requirements for a Growing Biotech
What “Validated System” Actually Means
For a computerized system supporting GxP processes, validation is the documented evidence that the system consistently does what it’s intended to do, and that it was implemented, tested, and is maintained in a controlled, repeatable way. For an eQMS, that typically includes a validation plan, documented user and functional requirements, risk-based testing (installation qualification, operational qualification, and performance qualification, depending on the risk assessment), traceability between requirements and test cases, and a summary report tying the evidence together. Vendors who support computer system validation (CSV) or computer software assurance (CSA) approaches, and who provide validation documentation packages or accelerators, can meaningfully reduce the burden this places on a biotech’s quality and IT teams.
Validation as the Company Scales
Validation isn’t a one-time event. Configuration changes, new modules, workflow updates, and system upgrades all need to be assessed for validation impact, and revalidated or retested where the risk assessment calls for it. A growing biotech that adds a new module or expands the system to a new site should expect to repeat a scoped version of this exercise. An eQMS that provides built-in validation documentation support, change-impact assessment tools, and release testing protocols makes this an ongoing, manageable discipline rather than a recurring fire drill. This is also where cloud-based, vendor-validated infrastructure can reduce the validation burden on the customer side, provided the vendor’s own validation evidence and change control practices are transparent and auditable.
Change Management Requirements in a Biotech QMS
Change is constant in biotech, which makes change management one of the more heavily scrutinized areas during audits and inspections. A QMS needs to handle at least three distinct categories of change.
Process Changes
When a manufacturing process, analytical method, or operating procedure changes, the QMS should route that change through a documented change control workflow that captures the reason for the change, an impact assessment, required approvals, and implementation and verification steps. The system should maintain a complete history of what changed, when, and why, tied back to the affected documents, records, and training requirements.
System Changes
Changes to the eQMS itself (configuration updates, new workflows, integrations, or upgrades) need their own change control process, distinct from process changes but following the same discipline. This is where the link to validation becomes direct. A system change that affects a validated function should trigger a documented assessment of whether revalidation or targeted retesting is needed before the change goes live.
Document Control
Version control is the backbone of change management. The QMS should enforce controlled authoring, review, and approval cycles, maintain a clear and unambiguous document history, prevent use of outdated or superseded versions, and tie document revisions to the training records of everyone who needs to be retrained as a result. For a biotech revising SOPs frequently as processes mature, this document control discipline is what keeps the paper trail defensible during an inspection.
Data Integrity Requirements and ALCOA+ Principles
Data integrity sits underneath every other requirement on this list, because none of the validation or change management evidence matters if the underlying data can’t be trusted. Regulators and auditors generally assess data integrity against the ALCOA+ principles, meaning data should be Attributable, Legible, Contemporaneous, Original, Accurate, and additionally Complete, Consistent, Enduring, and Available.
Audit Trails
An eQMS should maintain a complete, tamper-evident audit trail for every GxP-relevant record, capturing who made a change, what was changed, when it happened, and, where applicable, why. Audit trails need to be readily reviewable, not buried in a format that requires IT intervention to extract or interpret. This is one of the most common inspection focus areas, and a system that can produce a clear audit trail on demand removes a significant source of inspection risk.
Access Controls and Electronic Signatures
Role-based access controls limit who can create, review, approve, or modify records, which supports both data integrity and accountability. Electronic signature capture, applied in a way consistent with 21 CFR Part 11 requirements, should bind the signature to the specific record and action, with the meaning of the signature (such as “reviewed” or “approved”) clearly indicated. Together, access controls and e-signatures make it possible to demonstrate exactly who was responsible for each decision in the quality record, which is central to defending both data integrity and the overall quality system during an audit.
How an eQMS Addresses Each of These Areas Concretely
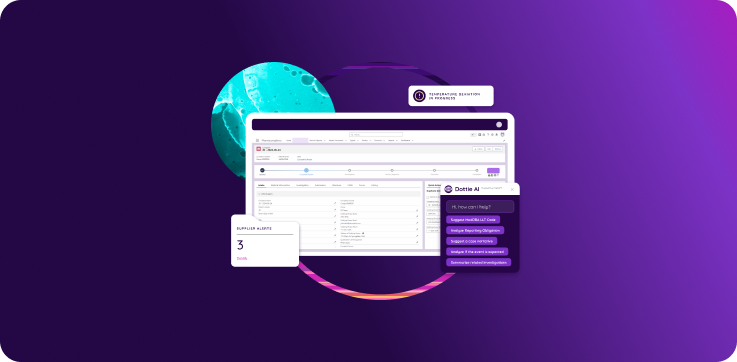
The common thread across growth, validation, change management, and data integrity is that they all depend on structure that holds up under pressure and under growth. A well-built eQMS supports this by combining configurable, scalable workflows with built-in validation documentation support, structured change control for both processes and the system itself, enforced document version control, and audit trails and access controls that satisfy ALCOA+ expectations without requiring manual reconstruction after the fact.
None of this replaces a biotech’s own quality judgment or its SOPs. What a capable QMS does is give the quality team a system that captures evidence consistently, scales with the business instead of against it, and holds up when an inspector asks to see exactly how a decision, a change, or a record came to be.
Choosing a QMS early with these requirements in mind, rather than retrofitting them in after a growth spurt or an inspection finding, tends to be the difference between a quality system that supports the business and one that the business has to work around.
Request a demo to see biotech-ready QMS capabilities.